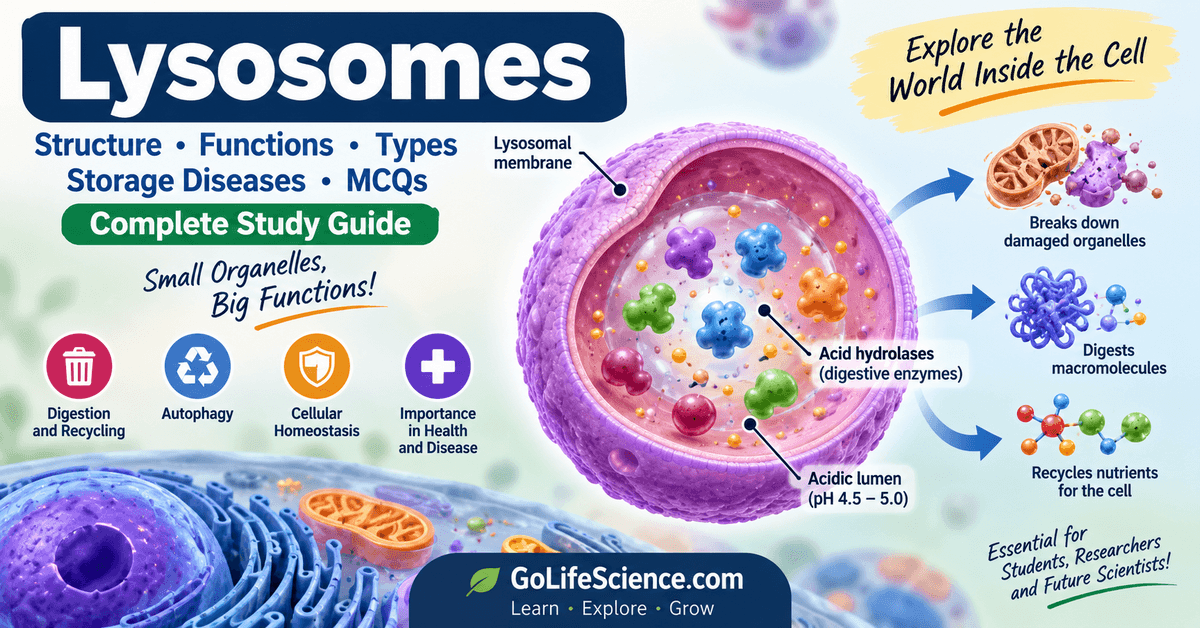
Lysosomes are membrane-bound organelles found in eukaryotic cells that play a vital role in the digestion, degradation, and recycling of cellular materials. Often described as the cell’s recycling centers, lysosomes contain a variety of acid hydrolases that break down proteins, lipids, carbohydrates, nucleic acids, and other unwanted cellular components.
A lysosome has a single membrane surrounding an acidic internal compartment, where its digestive enzymes work efficiently. The acidic environment is maintained mainly by V-type H⁺-ATPase proton pumps in the lysosomal membrane. Lysosomes receive materials through processes such as endocytosis, phagocytosis, and autophagy, digest them, and return useful breakdown products to the cell for reuse.
In addition to intracellular digestion, lysosomes are involved in autophagy, cellular recycling, nutrient sensing, signaling, and plasma membrane repair. Their formation involves the endomembrane system, including the rough endoplasmic reticulum, Golgi apparatus, and endosomal compartments.
In this guide, you will learn about lysosome structure, lysosomal enzymes, types of lysosomes, lysosome formation and biogenesis, major functions, autophagy, lysosomal storage diseases, and important exam points. This makes the article useful for students studying cell biology, biochemistry, biotechnology, medicine, and competitive examinations.
Quick Facts About Lysosomes
| Feature | Description |
|---|---|
| Organelle | Lysosome |
| Membrane | Single membrane-bound organelle |
| Location | Mainly found in the cytoplasm of eukaryotic cells |
| Main contents | Acid hydrolases and other lysosomal proteins |
| Internal environment | Acidic |
| Approximate pH | About 4.5–5.0 |
| Main function | Digestion and recycling of cellular materials |
| Major processes | Endocytosis, phagocytosis, and autophagy |
| Important proton pump | V-type H⁺-ATPase |
| Major enzyme groups | Proteases, lipases, nucleases, glycosidases, phosphatases, and sulfatases |
| Enzyme targeting | Many lysosomal enzymes are sorted through the mannose-6-phosphate pathway |
| Major cellular role | Degradation, recycling, cellular quality control, and metabolic regulation |
| Associated diseases | Lysosomal storage diseases |
| Discovery | Lysosomes were identified through the work of Christian de Duve and colleagues |
Lysosomes are dynamic organelles rather than simple waste-disposal compartments.
Their acidic lumen allows many acid hydrolases to break down biological molecules efficiently, while transport systems return useful degradation products to the cytoplasm.
Lysosomes also communicate with other cellular compartments and participate in processes such as autophagy, nutrient sensing, and membrane repair.
What Are Lysosomes?
Lysosomes are membrane-bound organelles that contain digestive enzymes called acid hydrolases.
They are responsible for breaking down and recycling cellular materials, including proteins, lipids, carbohydrates, nucleic acids, damaged organelles, and material taken into the cell.
Lysosomes are mainly found in eukaryotic cells, particularly animal cells, and form an important part of the cell’s endomembrane system. Each lysosome is surrounded by a single membrane that separates its acidic interior from the cytoplasm.
The inside of a lysosome is maintained at an acidic pH of approximately 4.5–5.0. This acidic environment is essential because many lysosomal enzymes work most efficiently under acidic conditions.
A V-type H⁺-ATPase (V-ATPase) in the lysosomal membrane uses energy from ATP to pump protons into the lysosomal lumen and maintain this acidic environment.
Lysosomes receive materials from several cellular pathways. For example, substances taken up through endocytosis can be delivered to lysosomal compartments for degradation. In phagocytosis, lysosomes help digest material engulfed by specialized cells.
During autophagy, damaged organelles and other cellular components are enclosed within autophagosomes, which subsequently fuse with lysosomal compartments for degradation.
After digestion, many useful products such as amino acids, sugars, fatty acids, and other small molecules can be transported back into the cytoplasm and reused by the cell.
Therefore, lysosomes are important not only for cellular digestion but also for recycling, cellular quality control, metabolism, and adaptation to changing conditions.
What Is the Main Function of a Lysosome?
The main function of a lysosome is to digest and recycle cellular materials using acid hydrolases. Lysosomes help remove damaged cellular components and break down materials delivered through endocytosis, phagocytosis, and autophagy.
When lysosomal enzymes or lysosomal trafficking pathways are defective, substances that normally would be degraded can accumulate inside cells. Such defects can cause a group of inherited disorders known as lysosomal storage diseases.
Key Points About Lysosomes
- Lysosomes are membrane-bound organelles.
- They contain numerous acid hydrolases.
- Their internal environment is acidic, generally around pH 4.5–5.0.
- V-ATPase helps maintain the acidic lysosomal lumen.
- They digest and recycle cellular materials.
- They participate in endocytosis, phagocytosis, and autophagy.
- Lysosomal dysfunction can lead to lysosomal storage diseases.
Discovery of Lysosomes
Lysosomes were identified in the 1950s through the work of Belgian biochemist Christian de Duve and his colleagues. Their discovery emerged from studies of cellular enzymes and subcellular compartments using biochemical fractionation and microscopy.
Christian de Duve was investigating enzymes such as acid phosphatase, an enzyme that showed greater activity when cell fractions were disrupted. This observation suggested that the enzymes were enclosed within membrane-bound structures rather than freely distributed throughout the cell.
These enzyme-containing structures were later recognized as a distinct cellular compartment and named lysosomes, derived from Greek words associated with loosening or breaking down and bodies.
Christian de Duve and Lysosome Discovery
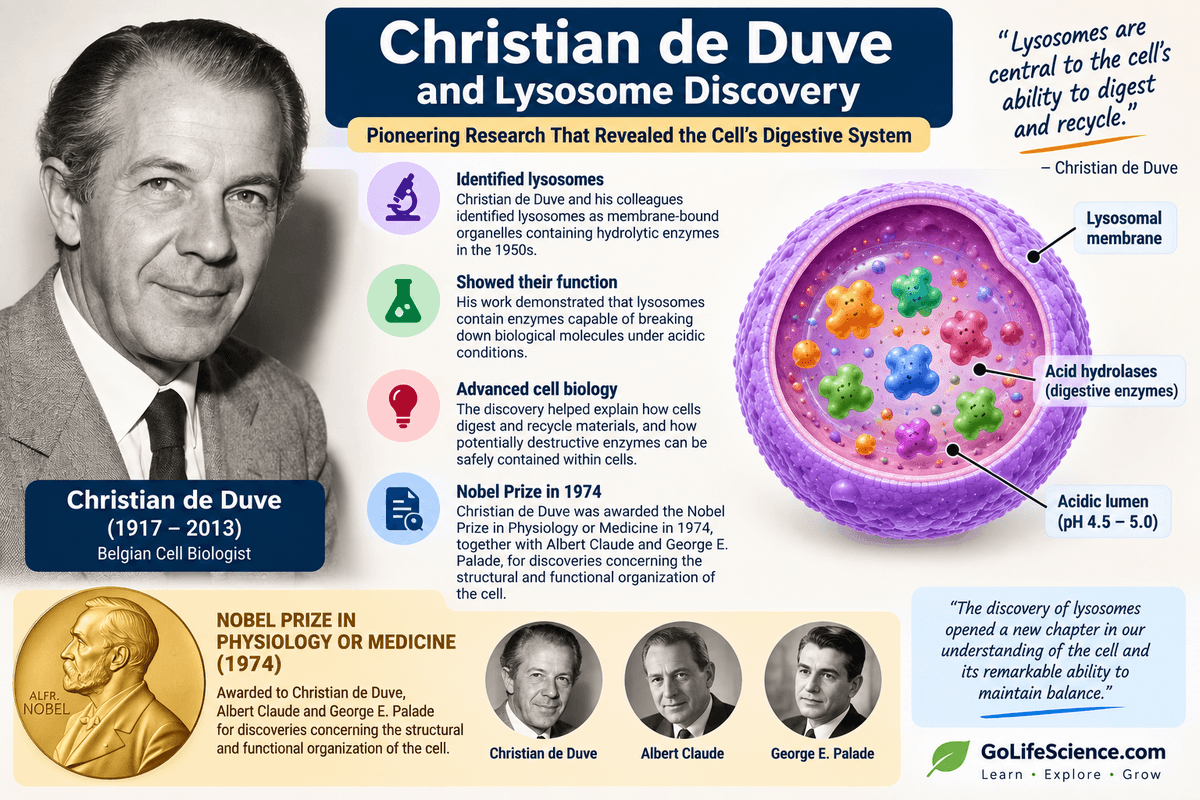
Christian de Duve played a central role in establishing lysosomes as important cellular organelles. His work demonstrated that lysosomes contain hydrolytic enzymes capable of breaking down biological molecules under acidic conditions.
The discovery significantly advanced the understanding of how cells digest and recycle materials. It also helped explain how potentially destructive digestive enzymes can be safely contained within cells.
In 1974, Christian de Duve was awarded the Nobel Prize in Physiology or Medicine, together with Albert Claude and George E. Palade, for discoveries concerning the structural and functional organization of the cell.
Why Was the Discovery of Lysosomes Important?
The discovery of lysosomes provided an explanation for several important cellular processes, including:
- How cells degrade unwanted or damaged materials.
- How digestive enzymes are compartmentalized within cells.
- How cellular components can be broken down and recycled.
- How intracellular digestion is coordinated with other membrane-bound compartments.
- How defects in lysosomal enzymes can cause storage diseases.
Today, lysosomes are understood as dynamic organelles involved not only in intracellular digestion but also in autophagy, recycling, nutrient sensing, signaling, and cellular homeostasis.
Key Point for Exams: Lysosomes were discovered in the 1950s through the work of Christian de Duve and colleagues. Christian de Duve is therefore commonly associated with the discovery of lysosomes in biology and biochemistry textbooks.
Lysosome Structure
A lysosome is a small, membrane-bound organelle with a specialized internal environment that allows it to digest and recycle cellular materials. Although lysosomes vary in size, shape, and composition depending on the cell type and physiological condition, they share several important structural features.
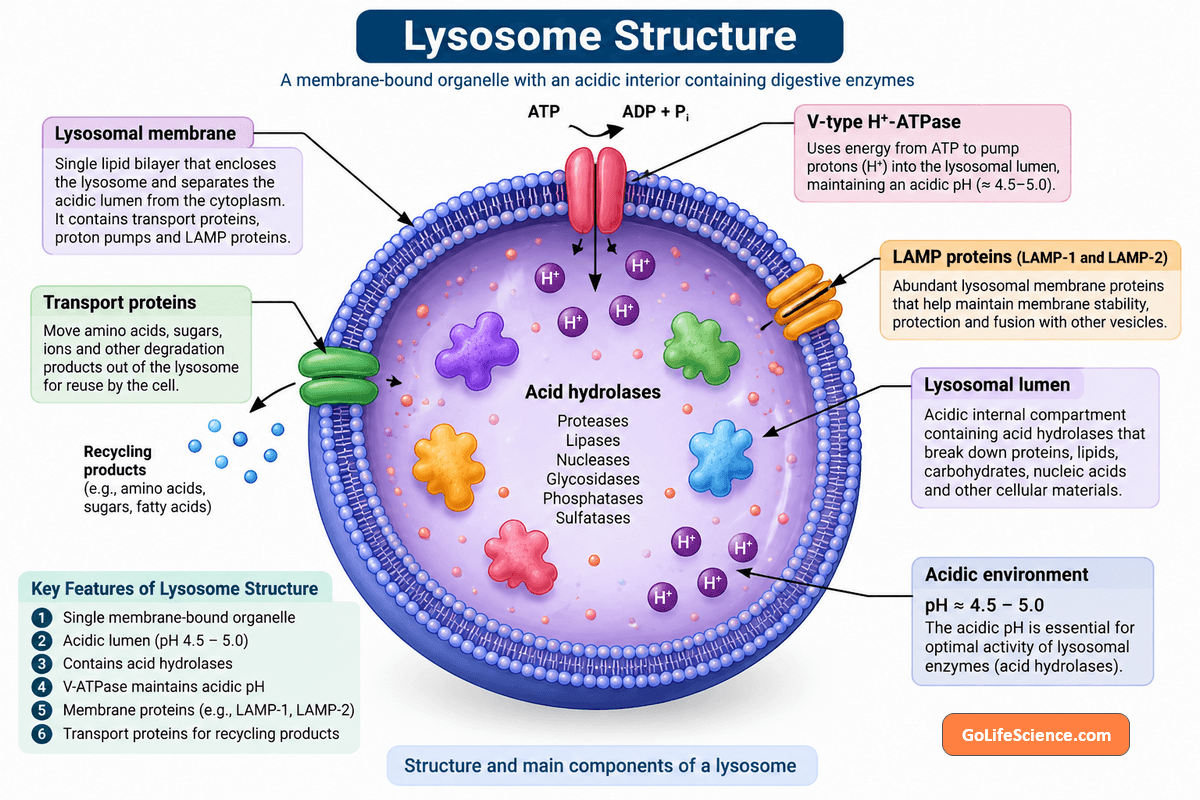
A typical lysosome consists of a single limiting membrane surrounding an acidic lumen that contains numerous digestive enzymes called acid hydrolases. The membrane also contains proteins responsible for transporting molecules, maintaining the acidic environment, protecting the membrane, and communicating with other cellular compartments.
1. Lysosomal Membrane
The lysosomal membrane is a single lipid bilayer that separates the acidic lysosomal lumen from the relatively neutral cytoplasm.
This membrane is important because it:
- Keeps lysosomal digestive enzymes contained within the organelle.
- Protects the cytoplasm from uncontrolled enzymatic degradation.
- Contains proton pumps that maintain lysosomal acidity.
- Contains transport proteins that move degradation products across the membrane.
- Helps lysosomes interact and fuse with endosomes, autophagosomes, and other vesicles.
Lysosomal membrane proteins also contribute to the stability and function of the organelle. Two important lysosomal-associated membrane proteins are LAMP-1 and LAMP-2, which are abundant components of the lysosomal membrane.
2. Lysosomal Lumen
The lysosomal lumen is the internal compartment enclosed by the lysosomal membrane. It contains a large collection of hydrolytic enzymes that digest biological molecules delivered to the lysosome.
These enzymes include
- Proteases
- Lipases
- Nucleases
- Glycosidases
- Phosphatases
- Sulfatases
The lumen provides the chemical conditions required for these enzymes to function efficiently.
3. Acidic pH of Lysosomes
One of the most important characteristics of lysosomes is their acidic internal environment. The lysosomal lumen generally has a pH of approximately 4.5–5.0, which is considerably more acidic than the surrounding cytoplasm.
This acidic environment is essential because many lysosomal enzymes, known as acid hydrolases, have their highest activity under acidic conditions.
Maintaining this pH also provides an additional level of protection for the cell. If lysosomal enzymes escape into the cytoplasm, many of them are less active at the near-neutral cytoplasmic pH.
4. V-Type H⁺-ATPase Proton Pump
The acidic environment inside lysosomes is maintained mainly by a V-type H⁺-ATPase (V-ATPase) located in the lysosomal membrane.
The V-ATPase uses energy from ATP to transport hydrogen ions (H⁺) from the cytoplasm into the lysosomal lumen.
The process can be summarized as
ATP energy → V-ATPase → H⁺ transported into lysosome → acidic lumen
Maintaining the proton gradient is essential for lysosomal enzyme activity and normal lysosomal function.
5. Lysosomal Membrane Proteins
The lysosomal membrane contains several groups of proteins with specialized functions.
- LAMP-1 and LAMP-2: These are abundant lysosomal membrane proteins that contribute to membrane organization and protection.
- V-ATPase: Maintains the acidic environment by transporting protons into the lysosomal lumen.
- Transport proteins: Move amino acids, sugars, ions, and other products generated during degradation out of the lysosome for reuse by the cell.
- Fusion and trafficking proteins: Help lysosomes interact with endosomes, autophagosomes, and other cellular compartments.
Summary of Lysosome Structure
The basic organization of a lysosome can therefore be represented as
Lysosomal membrane → acidic lumen → acid hydrolases → degradation of cellular materials → transport of useful products back to the cytoplasm
The structure of the lysosome is closely connected to its function. Its protective membrane, acidic lumen, digestive enzymes, proton pumps, and transport systems work together to allow the cell to safely degrade and recycle materials.
Lysosome Structure Diagram
The following lysosome structure diagram shows the major components of a typical lysosome, including its single membrane, acidic lumen, digestive enzymes, proton pump, membrane proteins, and transport proteins.
Major Parts Shown in the Lysosome Diagram
- Lysosomal membrane: A single lipid bilayer surrounds the lysosome and separates its acidic interior from the cytoplasm. It contains specialized proteins involved in transport, proton pumping, and membrane stability.
- Lysosomal lumen: The lumen is the internal compartment of the lysosome. It contains acid hydrolases and provides the acidic environment required for efficient enzymatic digestion.
- Acid hydrolases: These digestive enzymes break down proteins, lipids, carbohydrates, nucleic acids, and other cellular materials. Important enzyme groups include proteases, lipases, nucleases, glycosidases, phosphatases, and sulfatases.
- V-type H⁺-ATPase: This proton pump uses energy from ATP to transport hydrogen ions (H⁺) into the lysosomal lumen. It helps maintain the acidic lysosomal pH, generally around 4.5–5.0.
- LAMP proteins: Lysosome-associated membrane proteins such as LAMP-1 and LAMP-2 are abundant membrane proteins that contribute to lysosomal membrane organization and protection.
- Transport proteins: Transport proteins move useful products generated during digestion, such as amino acids, sugars, and fatty acids, across the lysosomal membrane so they can be reused by the cell.
How the Lysosome Structure Supports Its Function
The structure of a lysosome is closely related to its digestive and recycling functions. The membrane keeps the enzymes contained, the V-ATPase maintains the acidic environment, and the acid hydrolases break down cellular materials. Transport proteins then help return useful degradation products to the cytoplasm.
Together, these components allow lysosomes to function as important centers for cellular degradation, recycling, and quality control.
Related: Learn more about the [structure and functions of cell organelles] and how lysosomes interact with other components of the endomembrane system.
Lysosomal Enzymes
Lysosomes contain a large collection of digestive enzymes known as acid hydrolases. These enzymes break down different types of biological molecules under acidic conditions. They are essential for the lysosome’s role in intracellular digestion, cellular recycling, autophagy, and removal of damaged cellular components.
Most lysosomal enzymes are synthesized in the rough endoplasmic reticulum (RER) and subsequently processed and sorted through the Golgi apparatus before being delivered to lysosomal compartments.
Major Types of Lysosomal Enzymes
| Enzyme group | Main function | Major substrates |
|---|---|---|
| Proteases | Break down proteins | Proteins and peptides |
| Lipases | Digest lipids | Triglycerides and other lipids |
| Nucleases | Break down nucleic acids | DNA and RNA |
| Glycosidases | Break down carbohydrates | Glycoproteins, glycolipids, and polysaccharides |
| Phosphatases | Remove phosphate groups | Phosphorylated molecules |
| Sulfatases | Remove sulfate groups | Sulfated carbohydrates and lipids |
What Are Acid Hydrolases?
Acid hydrolases are hydrolytic enzymes that function efficiently under acidic conditions. The term “hydrolase” refers to enzymes that use water to break chemical bonds in their substrates.
The acidic environment inside lysosomes allows these enzymes to work efficiently. Different hydrolases act on different types of molecules, allowing lysosomes to digest a wide variety of cellular materials.
For example, proteases digest proteins, lipases break down lipids, and nucleases degrade DNA and RNA.
How Are Lysosomal Enzymes Targeted to Lysosomes?
Lysosomal enzymes follow a specialized trafficking pathway before reaching their destination.
The basic pathway is
Rough ER → Golgi apparatus → Mannose-6-phosphate tagging → Sorting → Endosomal pathway → Lysosome
Many lysosomal enzymes receive a mannose-6-phosphate (M6P) modification in the Golgi apparatus. M6P receptors recognize these tagged enzymes and help direct them toward endosomal compartments and ultimately the lysosomal system.
This targeting mechanism is important because lysosomal enzymes need to reach the correct cellular compartment to perform their digestive functions.
Why Are Lysosomal Enzymes Important?
Lysosomal enzymes help cells:
- Digest proteins, lipids, carbohydrates, and nucleic acids.
- Remove damaged or unwanted cellular components.
- Break down material taken up through endocytosis and phagocytosis.
- Digest cellular components delivered through autophagy.
- Recycle useful molecules for reuse by the cell.
- Maintain cellular homeostasis.
What Happens When Lysosomal Enzymes Are Defective?
A deficiency or dysfunction of a lysosomal enzyme can prevent a particular substance from being completely degraded. The undegraded material may progressively accumulate inside lysosomes and other cellular compartments.
Such defects can cause lysosomal storage diseases, including Gaucher disease, Tay-Sachs disease, Pompe disease, and Fabry disease.
Therefore, lysosomal enzymes are important not only for normal cellular digestion but also for understanding several inherited metabolic disorders.
Key Point for Exams: Lysosomes contain acid hydrolases that digest different classes of biological molecules under acidic conditions. Many lysosomal enzymes are synthesized in the rough ER, processed in the Golgi apparatus, and targeted to the lysosomal system through the mannose-6-phosphate pathway.
Lysosome Formation and Biogenesis
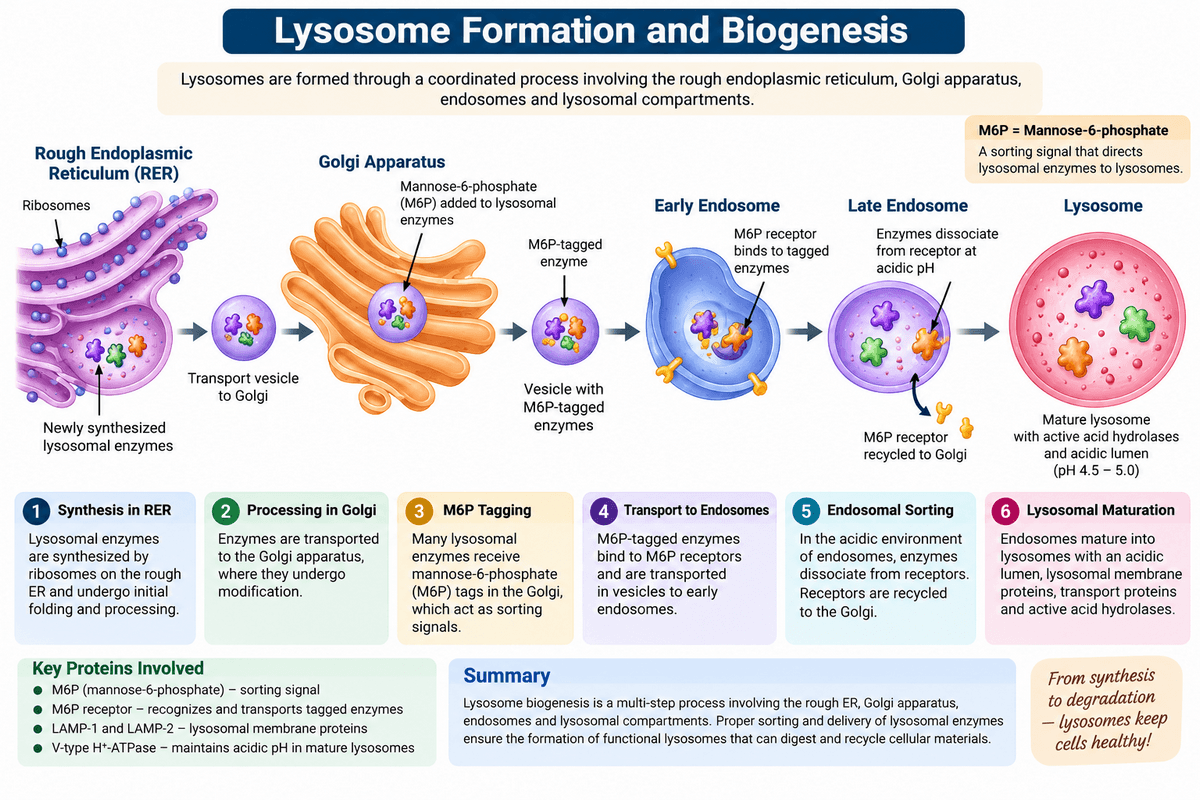
Lysosome formation is a coordinated process involving the rough endoplasmic reticulum (RER), Golgi apparatus, endosomes, and lysosomal compartments. Lysosomes are not simply produced as fully formed organelles by the Golgi apparatus. Instead, lysosomal proteins and enzymes move through the endomembrane system and are sorted into compartments that ultimately develop into functional lysosomes.
How Are Lysosomes Formed?
The formation and maturation of the lysosomal system can be simplified into the following pathway:
Rough ER → Golgi apparatus → Mannose-6-phosphate tagging → Endosomal sorting → Lysosomal maturation
Step 1: Synthesis of Lysosomal Enzymes in the Rough ER
Many lysosomal enzymes are synthesized by ribosomes attached to the rough endoplasmic reticulum. These enzymes enter the ER, where they undergo initial folding and processing before being transported toward the Golgi apparatus.
Step 2: Processing in the Golgi Apparatus
The newly synthesized enzymes are transported from the ER to the Golgi apparatus. During their passage through the Golgi, they undergo additional modifications and are prepared for delivery to the lysosomal system.
One of the most important targeting signals is mannose-6-phosphate (M6P).
Step 3: Mannose-6-Phosphate Tagging
Many lysosomal enzymes receive an M6P modification that acts as a molecular address label.
M6P receptors in the appropriate Golgi/endosomal trafficking system recognize these enzymes and help direct them toward endosomal compartments rather than allowing them to follow pathways intended for secretion or other cellular destinations.
Step 4: Transport to Endosomal Compartments
M6P-tagged lysosomal enzymes are transported in vesicles to endosomes. The acidic environment of endosomal compartments promotes the separation of many enzymes from their M6P receptors.
The receptors can be recycled, while the enzymes continue toward lysosomal compartments.
Step 5: Lysosomal Maturation
Endosomal compartments undergo maturation and progressively acquire the characteristics of degradative lysosomes, including:
- An increasingly acidic lumen.
- Lysosomal membrane proteins.
- Acid hydrolases.
- Transport proteins.
- Increased capacity for degradation.
The resulting lysosomal system can then receive material from endocytosis, phagocytosis, and autophagy for degradation.
Role of the Golgi Apparatus in Lysosome Formation
The Golgi apparatus plays an important role in processing and sorting many lysosomal enzymes, but it is more accurate to describe lysosome biogenesis as a coordinated process involving the ER, Golgi, endosomes, and lysosomal compartments.
The Golgi helps modify and sort lysosomal enzymes, while the endosomal system participates in their delivery and the maturation of degradative compartments.
Why Is Lysosome Biogenesis Important?
Proper lysosome formation ensures that digestive enzymes reach the correct cellular compartment and that lysosomes contain the proteins required for degradation and recycling.
Defects in lysosomal enzyme targeting, trafficking, or lysosomal maturation can interfere with cellular degradation and contribute to disease.
Key Point for Exams: Lysosome biogenesis involves the endomembrane system. Many lysosomal enzymes are synthesized in the rough ER, processed and sorted through the Golgi apparatus, targeted using mannose-6-phosphate signals, and delivered through endosomal compartments that contribute to lysosomal maturation.
Types of Lysosomes
Lysosomes have traditionally been classified according to their stage of formation, contents, and interaction with materials to be degraded. In basic cell biology textbooks, four commonly discussed forms are primary lysosomes, secondary lysosomes, autolysosomes, and residual bodies.
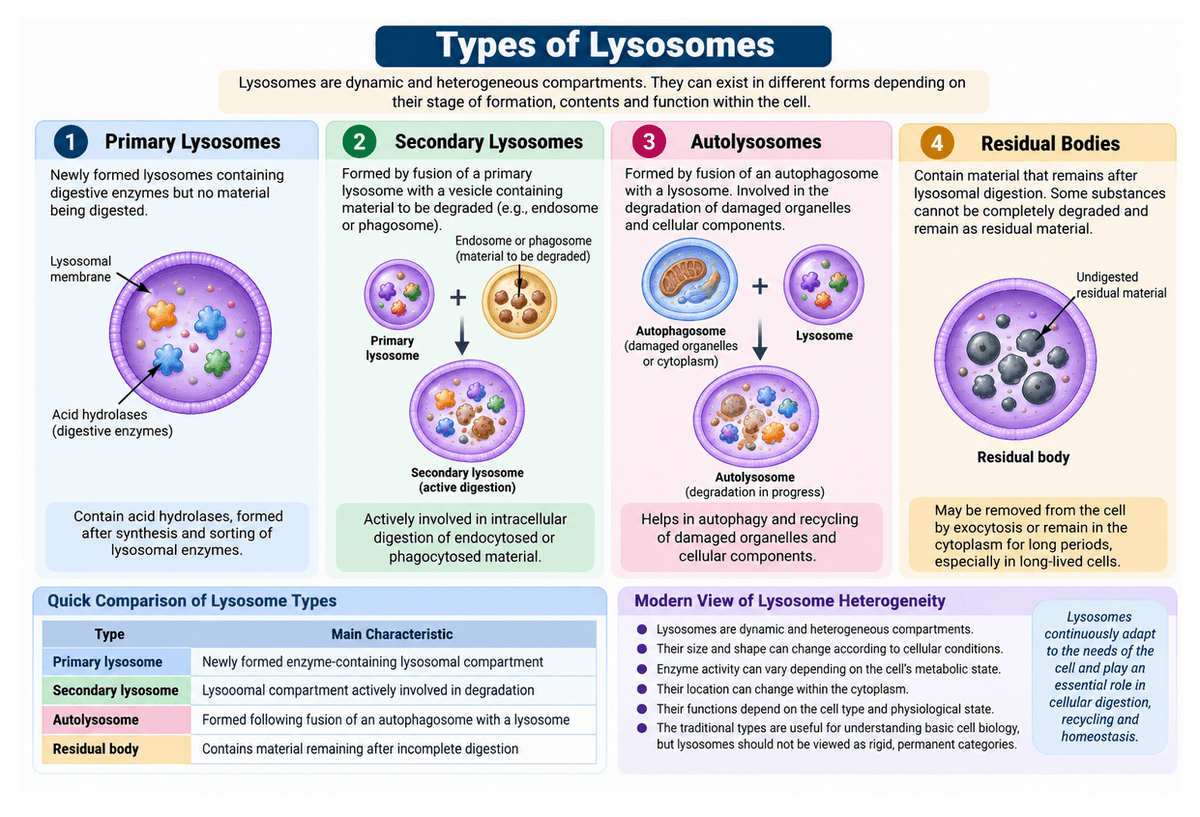
However, modern cell biology provides a more dynamic view. Lysosomes are not fixed structures that always fit into clearly separated categories. Their size, shape, location, enzyme activity, and functions can change according to the cell type and its physiological state.
1. Primary Lysosomes
Primary lysosomes are newly formed lysosomal compartments containing hydrolytic enzymes but little or no material undergoing active digestion. They are associated with the lysosomal system after lysosomal enzymes have been synthesized, processed, and delivered through the endomembrane pathway.
Primary lysosomes can subsequently interact with vesicles or compartments containing materials that need to be degraded.
2. Secondary Lysosomes
Secondary lysosomes are traditionally described as lysosomes formed when a primary lysosome fuses with a vesicle or compartment containing material to be digested.
For example, a lysosome may fuse with an endosomal or phagocytic compartment containing extracellular material. The acid hydrolases within the lysosomal compartment then break down the material into smaller molecules that can be reused by the cell.
Secondary lysosomes are therefore closely associated with active intracellular digestion.
3. Autolysosomes
An autolysosome is a degradative compartment formed when an autophagosome fuses with a lysosomal compartment.
During autophagy, damaged organelles or portions of the cytoplasm are enclosed within a double-membrane structure called an autophagosome. The autophagosome subsequently fuses with a lysosome, allowing lysosomal enzymes to degrade its contents.
The resulting degradation products can then be released for recycling and reuse by the cell.
4. Residual Bodies
A residual body is a compartment containing material that remains after lysosomal digestion. Some cellular materials cannot be completely degraded and may therefore remain as residual material.
Residual bodies may be removed from the cell through exocytosis or, in some long-lived cells, remain within the cytoplasm for extended periods.
Modern View of Lysosome Heterogeneity
Modern research shows that lysosomes are dynamic and heterogeneous organelles rather than a collection of rigidly defined types.
Their:
- Size and shape can change according to cellular conditions.
- Enzyme activity can vary depending on the cell’s metabolic state.
- Location can change within the cytoplasm.
- Interactions with other organelles can change during processes such as autophagy and nutrient deprivation.
- Functions can differ between cell types and physiological conditions.
Therefore, the traditional terms primary lysosome, secondary lysosome, autolysosome, and residual body are useful for understanding basic cell biology and examination concepts, but they should not be interpreted as completely separate, permanent lysosome categories.
Quick Comparison of Lysosome Types
| Type | Main characteristic |
|---|---|
| Primary lysosome | Newly formed enzyme-containing lysosomal compartment |
| Secondary lysosome | Lysosomal compartment actively involved in degradation |
| Autolysosome | Formed following fusion of an autophagosome with a lysosome |
| Residual body | Contains material remaining after incomplete digestion |
Key point: Lysosome types and functions are closely related to the cellular pathway delivering material for degradation. Modern cell biology emphasizes that lysosomes continuously change their structure and activity according to cellular needs.
Functions of Lysosomes
Lysosomes perform several important functions that help cells digest, recycle, and maintain cellular components.
Their acid hydrolases can break down proteins, lipids, carbohydrates, nucleic acids, and other materials delivered to the lysosomal system.
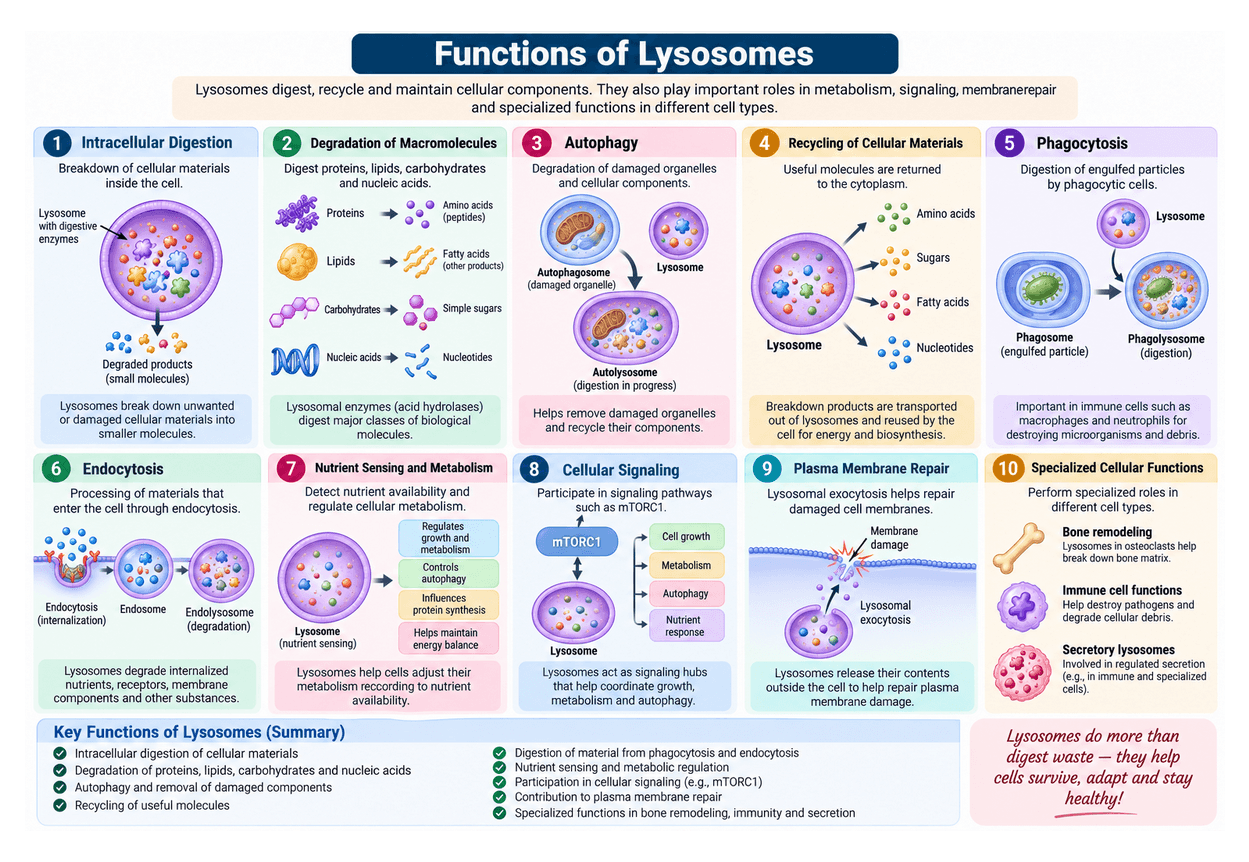
In addition to intracellular digestion, modern cell biology has shown that lysosomes participate in autophagy, nutrient sensing, metabolism, cellular signaling, membrane repair, and specialized functions in different cell types.
1. Intracellular Digestion
One of the primary functions of lysosomes is intracellular digestion. Lysosomes receive materials from different cellular pathways and use acid hydrolases to break them down into smaller molecules.
The material may originate from inside the cell, such as damaged organelles, or from outside the cell through endocytosis and phagocytosis. After degradation, useful products can be transported out of the lysosome and reused by the cell.
2. Degradation of Macromolecules
Lysosomes contain different groups of hydrolytic enzymes, allowing them to digest several major classes of biological molecules.
- Proteins: Proteases break proteins into peptides and amino acids.
- Lipids: Lipases and related enzymes break down lipids into smaller components such as fatty acids and other molecules.
- Carbohydrates: Glycosidases degrade complex carbohydrates and carbohydrate-containing molecules.
- Nucleic acids: Nucleases break down DNA and RNA into smaller nucleotide components.
This broad enzymatic activity makes lysosomes important centers for cellular degradation and recycling.
3. Autophagy
Lysosomes play a central role in autophagy, a cellular quality-control process in which damaged or unnecessary cellular components are delivered to lysosomal compartments for degradation.
During macroautophagy, cellular material is enclosed within an autophagosome. The autophagosome then fuses with a lysosomal compartment to form an autolysosome, where lysosomal enzymes digest the contents.
Autophagy helps cells remove damaged organelles and recycle their molecular components, particularly during cellular stress or nutrient limitation.
4. Recycling of Cellular Materials
Lysosomal digestion does not simply destroy cellular material. An important lysosomal function is to recover useful molecules.
Following degradation, products such as:
- Amino acids
- Sugars
- Fatty acids
- Nucleotides
- Other small metabolites
can be transported from lysosomal compartments into the cytoplasm. The cell can then reuse these molecules for energy production, biosynthesis, and other metabolic processes.
Therefore, lysosomes act as important cellular recycling centers.
5. Phagocytosis
Lysosomes also participate in phagocytosis, a process in which specialized cells engulf large particles such as microorganisms, cellular debris, or foreign material.
After a particle is engulfed, it becomes enclosed within a membrane-bound phagosome. The phagosome can subsequently interact and fuse with lysosomal compartments, exposing the engulfed material to acid hydrolases and other degradative mechanisms.
This function is particularly important in immune cells such as macrophages and neutrophils.
6. Endocytosis
Lysosomes participate in the processing of materials that enter cells through endocytosis. During endocytosis, portions of the plasma membrane surround extracellular molecules or membrane components and form intracellular vesicles.
These vesicles move through the endosomal system. Material that requires degradation can eventually be delivered to lysosomal compartments, where enzymes break it down.
This pathway allows cells to process extracellular nutrients, receptors, membrane components, and other internalized substances.
7. Nutrient Sensing and Metabolism
Modern research has established that lysosomes have functions beyond digestion. They act as important sites for nutrient sensing and metabolic regulation.
Lysosomes can detect changes in the availability of nutrients and communicate this information to signaling pathways that regulate processes such as:
- Protein synthesis
- Autophagy
- Cellular growth
- Energy metabolism
- Nutrient utilization
Through these activities, lysosomes help cells adjust their metabolism according to changing environmental and nutritional conditions.
8. Cellular Signaling
Lysosomes also participate in cellular signaling. Proteins and signaling complexes associated with lysosomal membranes can help regulate pathways involved in cell growth, metabolism, autophagy, and nutrient availability.
One important example is the relationship between lysosomes and the mTORC1 signaling pathway, which helps cells coordinate growth and metabolism with nutrient availability.
Thus, lysosomes function as more than digestive compartments; they also act as important signaling hubs.
9. Plasma Membrane Repair
Lysosomes can contribute to plasma membrane repair when the cell membrane is damaged.
In response to certain types of membrane injury, lysosomes can move toward the damaged region and undergo lysosomal exocytosis. This process releases lysosomal contents outside the cell and contributes to membrane repair mechanisms.
This function demonstrates how lysosomes can participate in cellular responses to physical damage.
10. Specialized Cellular Functions
Lysosomal compartments also perform specialized functions depending on the cell type.
- Bone remodeling: Lysosome-related degradative mechanisms contribute to the activity of osteoclasts, specialized cells involved in bone resorption.
- Immune cell functions: Lysosomal and lysosome-related compartments help immune cells degrade engulfed microorganisms and cellular debris.
- Secretory lysosomes: Some specialized cells contain secretory lysosomes that combine degradative functions with regulated secretion. These compartments are particularly important in certain immune and specialized cell types.
Summary of Lysosome Functions
The major functions of lysosomes can be summarized as
| Function | Main role |
|---|---|
| Intracellular digestion | Break down cellular materials |
| Macromolecule degradation | Digest proteins, lipids, carbohydrates, and nucleic acids |
| Autophagy | Remove and degrade damaged cellular components |
| Recycling | Return useful molecules to the cytoplasm |
| Phagocytosis | Digest engulfed particles and microorganisms |
| Endocytosis | Process internalized cellular and extracellular material |
| Nutrient sensing | Help regulate cellular metabolism |
| Cellular signaling | Participate in growth, metabolism, and autophagy pathways |
| Membrane repair | Contribute to repair of damaged plasma membranes |
| Specialized functions | Support bone remodeling, immunity, and regulated secretion |
Overall, the role of lysosomes in the cell extends far beyond simple waste disposal. Lysosomes are dynamic organelles that coordinate degradation, recycling, quality control, metabolism, signaling, and cellular adaptation, making them essential for maintaining cellular homeostasis.
Lysosomes and Autophagy
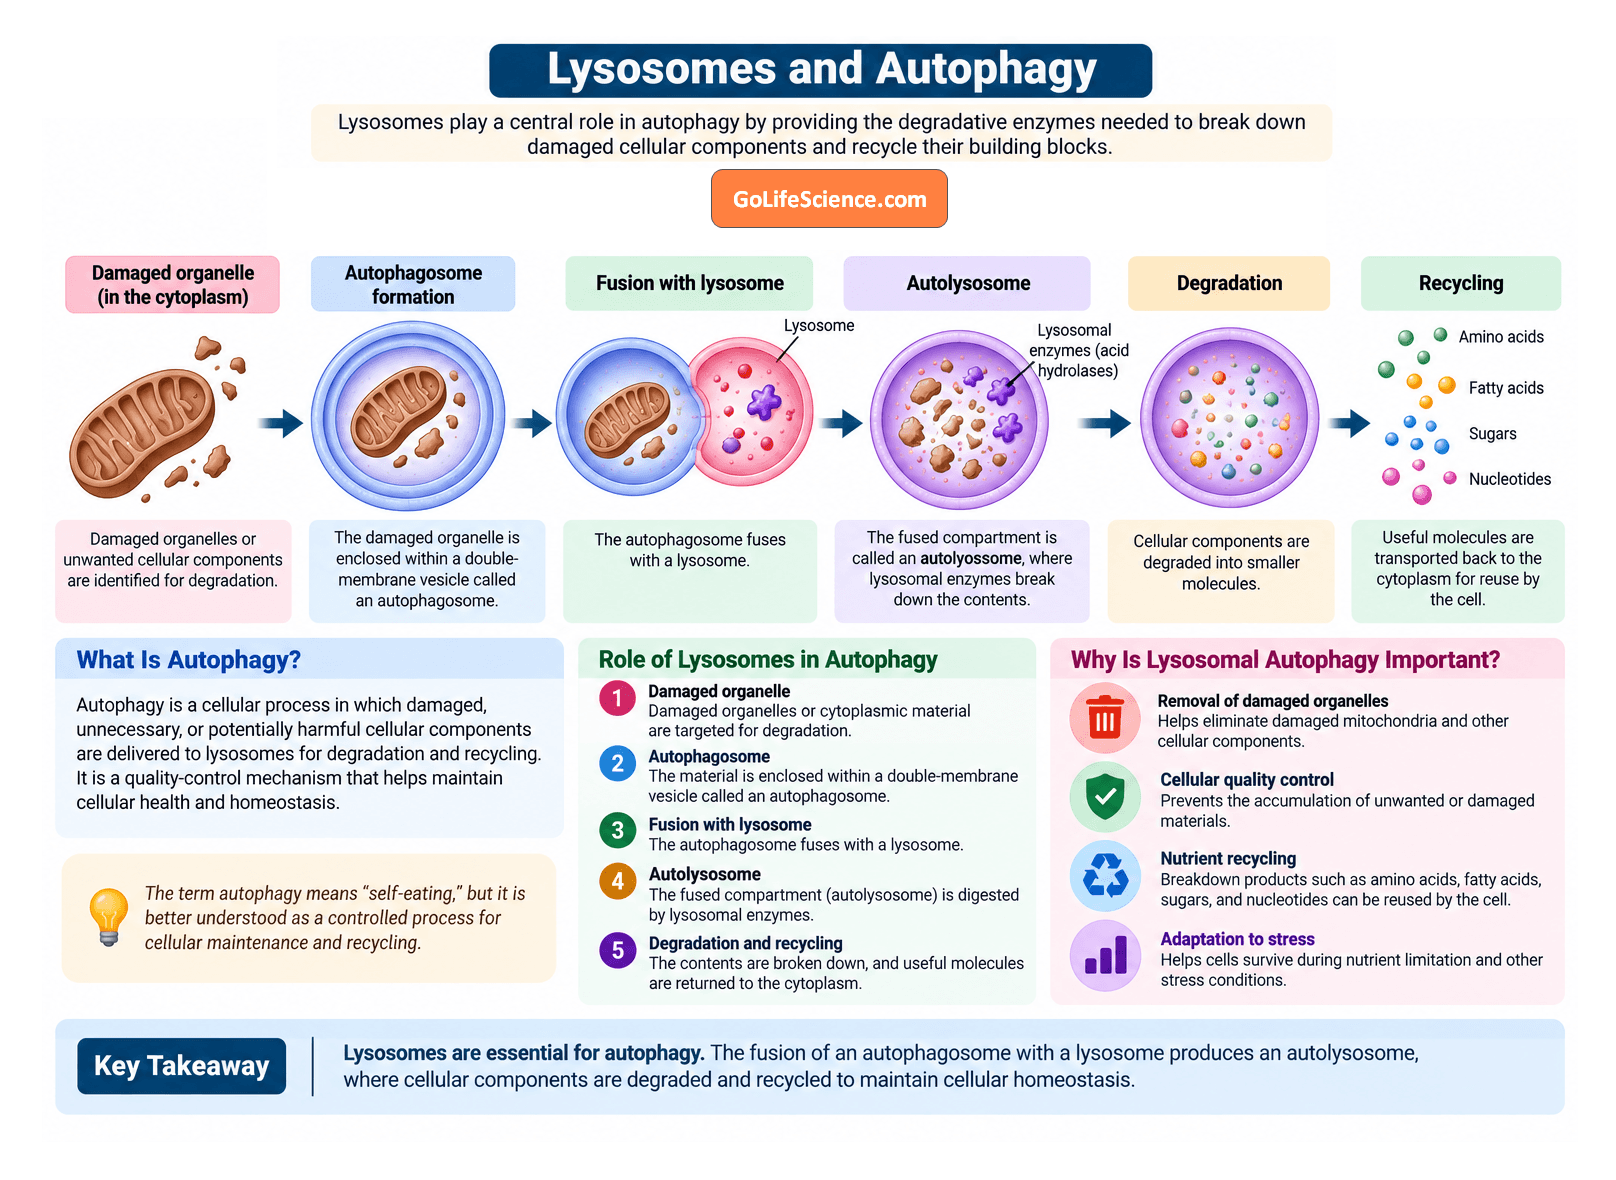
Lysosomes and autophagy are closely connected because lysosomes provide the main degradative machinery required to break down many cellular components targeted for autophagic recycling. Autophagy is an essential cellular quality-control process that helps cells remove damaged or unnecessary components and recover useful molecules from them.
What Is Autophagy?
Autophagy is a cellular process in which damaged, unnecessary, or potentially harmful cellular components are delivered to lysosomal compartments for degradation and recycling. The word “autophagy” means “self-eating,” but the process is better understood as a controlled mechanism for cellular maintenance and recycling.
During macroautophagy, damaged organelles or portions of cytoplasm are enclosed within a double-membrane structure called an autophagosome. The autophagosome then moves through the cell and eventually fuses with a lysosomal compartment.
How Lysosomes Participate in Autophagy
The relationship between lysosomes and autophagy can be summarized as:
Damaged organelle → Autophagosome → Fusion with lysosome → Autolysosome → Degradation → Recycling
After autophagosome–lysosome fusion, the resulting compartment is commonly called an autolysosome. Lysosomal acid hydrolases then break down the enclosed cellular material.
The resulting molecules, such as amino acids, fatty acids, sugars, and other metabolites, can be transported back into the cytoplasm and reused by the cell.
This process allows lysosomes to serve as the major degradative endpoint of macroautophagy.
Why Is Lysosomal Autophagy Important?
Lysosome-dependent autophagy performs several important functions:
- Removal of damaged organelles: It helps eliminate damaged mitochondria and other cellular components.
- Cellular quality control: It prevents the accumulation of unwanted or damaged material.
- Nutrient recycling: Breakdown products can be returned to the cytoplasm for reuse.
- Adaptation to stress: During conditions such as nutrient limitation, autophagy can help cells recycle internal resources and adapt to changing conditions.
Therefore, the role of lysosomes in autophagy extends beyond simple degradation. Lysosomes help maintain cellular homeostasis by connecting the removal of damaged components with the recycling of their molecular building blocks.
Key Point: Lysosome function in autophagy depends on the delivery of cellular material to lysosomal compartments, where it is degraded and recycled. The fusion of an autophagosome with a lysosome is a key step in this process and produces an autolysosome in which degradation takes place.
Why Are Lysosomes Called Suicidal Bags of the Cell?
Lysosomes are traditionally called the “suicidal bags of the cell” because they contain powerful digestive enzymes that can break down proteins, lipids, carbohydrates, nucleic acids, and other cellular materials.
The term is commonly used in school and college-level biology to emphasize the potentially destructive nature of lysosomal enzymes.
Why Are Lysosomes Called Suicidal Bags?
Lysosomes contain numerous acid hydrolases that normally remain enclosed within the lysosomal membrane.
If lysosomal membranes become severely damaged and digestive enzymes are released into the surrounding cellular environment, they can contribute to the degradation of cellular components.
Extensive enzyme release and lysosomal damage can therefore contribute to autolysis, the breakdown of a cell by its own enzymes.
The phrase “suicide bag of the cell” is therefore a useful traditional description for understanding the destructive potential of lysosomal enzymes.
However, it is important to recognize that the term is an oversimplification. Lysosomes do not normally function to destroy the cell.
Their primary roles include controlled degradation, recycling, autophagy, cellular quality control, nutrient regulation, and other essential cellular processes.
The lysosomal membrane and cellular regulatory mechanisms normally keep their digestive activities compartmentalized and controlled.
Exam Point
For biology examinations, remember:
Lysosomes are traditionally called the “suicidal bags of the cell” because they contain hydrolytic enzymes that can contribute to cellular self-digestion if released or improperly regulated.
This terminology is useful for exam preparation, but modern cell biology recognizes lysosomes as dynamic organelles essential for cellular digestion, recycling, autophagy, and homeostasis, rather than simply destructive structures.
Lysosomes in Plant and Animal Cells
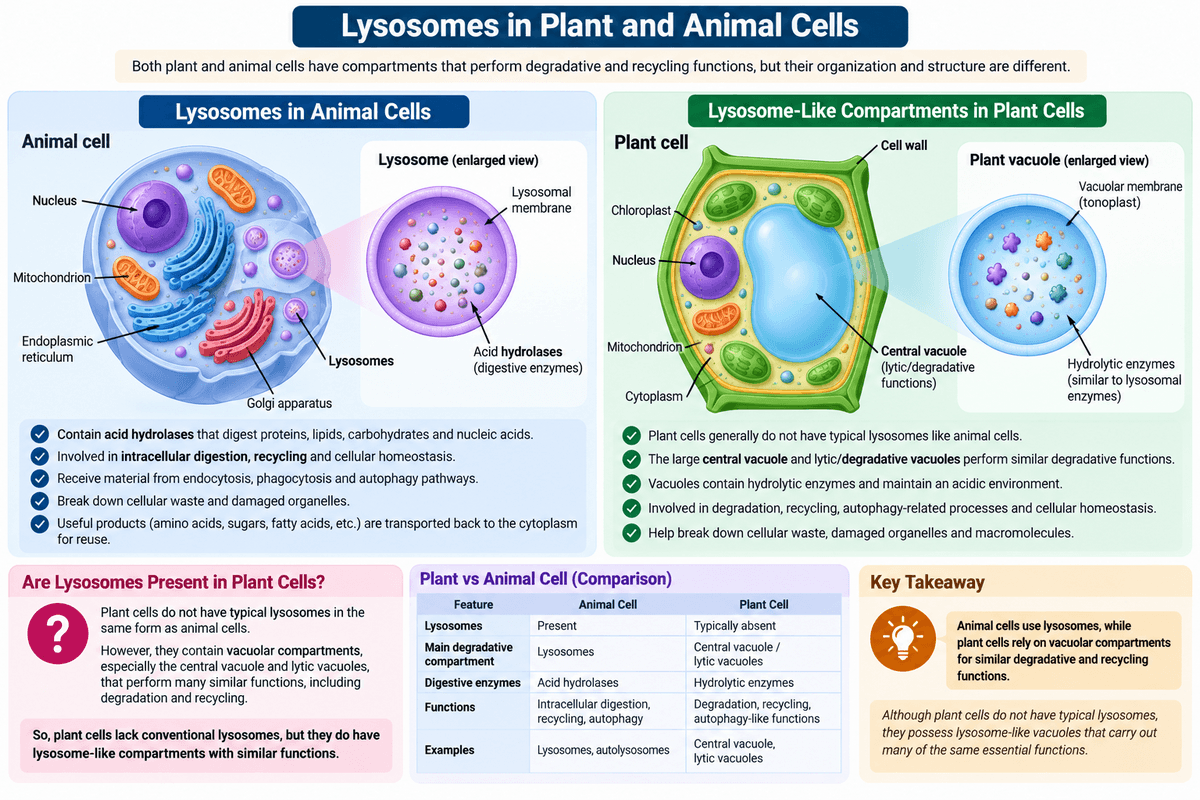
Lysosomes and lysosome-like compartments are involved in the degradation and recycling of cellular materials in both animals and plants, although their organization differs between the two types of cells.
Understanding this distinction helps avoid the common misconception that plants simply lack lysosomal functions.
1. Lysosomes in Animal Cells
Typical animal-cell lysosomes are membrane-bound organelles containing acid hydrolases. They receive materials from pathways such as endocytosis, phagocytosis, and autophagy and digest them under acidic conditions.
After degradation, useful products such as amino acids, sugars, fatty acids, and other small molecules can be transported back into the cytoplasm. In this way, lysosomes contribute to intracellular digestion, cellular recycling, and homeostasis.
Lysosomes are particularly important in specialized animal cells, including immune cells, where lysosome-related compartments help digest engulfed microorganisms and cellular debris.
2. Lysosome-Like Compartments in Plant Cells
Plant cells generally do not have the same type of typical lysosomes found in animal cells. Instead, many of their degradative and recycling functions are performed by vacuolar compartments, particularly the large central vacuole and specialized lytic or degradative vacuoles.
Plant vacuoles can contain hydrolytic enzymes and maintain an acidic environment, allowing them to break down cellular components and recycle their molecular building blocks. These functions overlap with several functions performed by lysosomes in animal cells.
3. Are Lysosomes Present in Plant Cells?
The most accurate answer is not in the typical animal-cell form. In basic biology, plant cells are commonly described as lacking conventional lysosomes because their degradative functions are largely performed by vacuoles.
However, saying that plant cells have no lysosomal functions is misleading. Plant vacuoles and lysosome-like compartments perform many comparable roles, including degradation, recycling, autophagy-related processes, and cellular homeostasis.
Key Point: Animal cells typically use lysosomes for intracellular digestion and recycling, while plant cells rely heavily on vacuolar compartments for similar degradative functions.
Lysosomal Storage Diseases
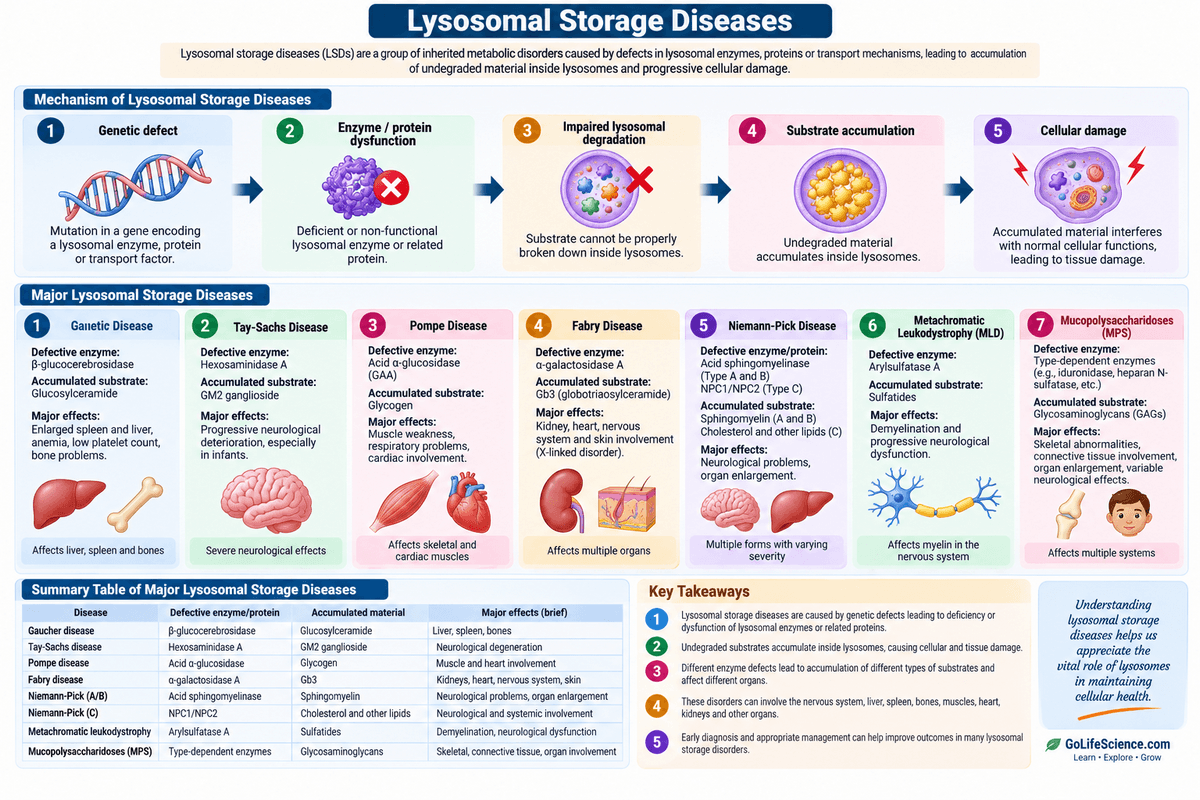
Lysosomal storage diseases (LSDs) are a group of inherited metabolic disorders in which lysosomal degradation or trafficking is impaired.
In many LSDs, a genetic defect causes deficiency or dysfunction of a lysosomal enzyme, leading to incomplete breakdown of a particular substrate.
The undegraded material then accumulates progressively inside cells and can interfere with normal cellular functions.
The basic mechanism can be summarized as
Genetic defect → Enzyme or protein dysfunction → Impaired lysosomal degradation → Substrate accumulation → Cellular dysfunction and tissue damage
Although enzyme deficiencies are common, some lysosomal storage disorders result from defects in lysosomal membrane proteins, transport proteins, activator proteins, or intracellular trafficking mechanisms.
1. Gaucher Disease
Gaucher disease is caused by deficient activity of β-glucocerebrosidase (glucosylceramidase). This prevents normal degradation of glucosylceramide, which accumulates particularly in cells of the monocyte-macrophage system.
Common effects can include enlargement of the spleen and liver, anemia, low platelet counts, and bone abnormalities or bone pain. Gaucher disease has several clinical forms with different patterns of neurological involvement.
2. Tay-Sachs Disease
Tay-Sachs disease is a GM2 gangliosidosis caused by deficiency of β-hexosaminidase A. The enzyme deficiency results in accumulation of GM2 ganglioside, particularly affecting nerve cells.
The severe infantile form can cause progressive neurological deterioration.
3. Pompe Disease
Pompe disease, also called glycogen storage disease type II, results from deficiency of acid α-glucosidase (GAA). This enzyme normally helps degrade lysosomal glycogen.
Consequently, glycogen accumulates, particularly in skeletal and cardiac muscle cells. Depending on the form, Pompe disease can cause muscle weakness, respiratory problems, and cardiac involvement.
4. Fabry Disease
Fabry disease is caused by deficient α-galactosidase A activity. This results in accumulation of the glycosphingolipid globotriaosylceramide (Gb3) and related metabolites.
Because Fabry disease is X-linked, its clinical manifestations can differ between affected males and females. The disease can involve multiple organs, including the kidneys, heart, nervous system, and skin.
5. Niemann-Pick Disease
Niemann-Pick diseases include several genetically and biochemically distinct disorders.
Niemann-Pick types A and B are associated with deficient acid sphingomyelinase, resulting in sphingomyelin accumulation. Type A generally has substantial neurological involvement, whereas type B is typically less neurologically severe.
Niemann-Pick type C is different: it involves defects in proteins responsible for intracellular lipid and cholesterol trafficking, particularly NPC1 or NPC2, leading to abnormal accumulation and distribution of cholesterol and other lipids.
6. Metachromatic Leukodystrophy
Metachromatic leukodystrophy (MLD) is commonly caused by deficiency of arylsulfatase A. This impairs the degradation of sulfatides, resulting in their accumulation, particularly in nervous tissue.
The progressive accumulation of sulfatides damages myelin and contributes to neurological dysfunction.
7. Mucopolysaccharidoses
Mucopolysaccharidoses (MPS) are a group of lysosomal storage disorders caused by deficiencies of different enzymes involved in the degradation of glycosaminoglycans (GAGs).
Depending on the specific MPS type, different GAGs such as heparan sulfate, dermatan sulfate, or keratan sulfate may accumulate. Clinical manifestations vary among the different disorders and can involve the skeletal system, connective tissues, respiratory system, heart, and nervous system.
Major Lysosomal Storage Diseases: Quick Comparison
| Disease | Defective enzyme/protein | Major accumulated material |
|---|---|---|
| Gaucher disease | β-glucocerebrosidase | Glucosylceramide |
| Tay-Sachs disease | Hexosaminidase A | GM2 ganglioside |
| Pompe disease | Acid α-glucosidase | Glycogen |
| Fabry disease | α-galactosidase A | Gb3 (globotriaosylceramide) |
| Niemann-Pick A/B | Acid sphingomyelinase | Sphingomyelin |
| Niemann-Pick C | NPC1/NPC2 proteins | Cholesterol and other lipids |
| Metachromatic leukodystrophy | Arylsulfatase A | Sulfatides |
| Mucopolysaccharidoses | Type-dependent enzymes | Glycosaminoglycans |
Key Point: The central feature of many lysosomal storage diseases is the progressive accumulation of substances that cells cannot properly degrade. The specific enzyme or protein affected determines the type of material stored and the tissues most severely affected. Diagnosis generally involves biochemical enzyme testing and/or molecular genetic testing, depending on the disorder.
Lysosomes – Important Points for Exams
For quick revision, remember these key points about lysosomes:
- Membrane-bound organelles: Lysosomes are surrounded by a single membrane.
- Acid hydrolases: They contain enzymes that digest proteins, lipids, carbohydrates, nucleic acids, and other materials.
- Acidic lumen: The lysosomal interior is maintained at an acidic pH, typically around 4.5–5.0.
- V-ATPase: V-type H⁺-ATPase pumps protons into the lysosomal lumen and maintains its acidity.
- Digestion and recycling: Lysosomes break down cellular materials and return useful molecules to the cell for reuse.
- Autophagy: They participate in the degradation and recycling of damaged organelles and cytoplasmic components.
- Endocytosis and phagocytosis: Lysosomes digest materials taken into cells through these pathways.
- Discovery: Christian de Duve and colleagues are associated with the discovery and characterization of lysosomes.
- Lysosomal storage diseases: Defects in lysosomal enzymes, proteins, or trafficking pathways can cause accumulation of undegraded materials.
- “Suicidal bags”: This is a traditional textbook term referring to the potential for lysosomal enzymes to contribute to cellular self-digestion when lysosomal integrity is severely disrupted.
Lysosomes – One-Minute Revision
Lysosomes are single-membrane organelles containing acid hydrolases. Their acidic lumen, maintained mainly by V-ATPase, allows these enzymes to digest proteins, lipids, carbohydrates, nucleic acids, and other cellular materials. Lysosomes receive material through endocytosis, phagocytosis, and autophagy, then degrade and recycle useful components. They are important for intracellular digestion, cellular quality control, nutrient recycling, and homeostasis. Christian de Duve is associated with their discovery. Defects in lysosomal function can cause lysosomal storage diseases.
Lysosomes MCQs (Test Your Knowledge)
Lysosomes MCQs
Test your knowledge of lysosome structure, functions, enzymes, autophagy and lysosomal storage diseases.
Your Result
Frequently Asked Questions About Lysosomes
What are lysosomes?
Lysosomes are single-membrane-bound organelles found mainly in animal cells. They contain acid hydrolases that digest proteins, lipids, carbohydrates, nucleic acids, and cellular waste. Lysosomes have an acidic internal environment that allows these enzymes to function efficiently. They are important for intracellular digestion, recycling, autophagy, and maintaining cellular homeostasis.
What is the main function of lysosomes?
The main function of lysosomes is to digest and recycle cellular materials. Lysosomal acid hydrolases break down proteins, lipids, carbohydrates, nucleic acids, damaged organelles, and material taken into the cell. The resulting smaller molecules can be transported back into the cytoplasm and reused for energy production, biosynthesis, or other cellular processes.
Why are lysosomes called suicidal bags?
Lysosomes are traditionally called “suicidal bags of the cell” because they contain powerful hydrolytic enzymes that can contribute to cellular self-digestion if released after severe lysosomal membrane damage. However, this term is an oversimplification. Under normal conditions, lysosomes are highly regulated organelles involved mainly in digestion, recycling, autophagy, and cellular quality control.
Who discovered lysosomes?
Christian de Duve and his colleagues are associated with the discovery of lysosomes in the 1950s. During biochemical studies of cellular enzymes, they identified a group of membrane-bound compartments containing hydrolytic enzymes. Christian de Duve later shared the 1974 Nobel Prize in Physiology or Medicine with Albert Claude and George E. Palade for discoveries concerning the structural and functional organization of the cell.
What enzymes are present in lysosomes?
Lysosomes contain many acid hydrolases, including proteases, lipases, nucleases, glycosidases, phosphatases, and sulfatases. These enzymes break down different types of biological molecules under acidic conditions. Proteases digest proteins, lipases break down lipids, nucleases act on nucleic acids, and glycosidases help degrade carbohydrates and complex glycoconjugates.
What is the pH of lysosomes?
The pH inside lysosomes is typically around 4.5–5.0, making the lysosomal lumen strongly acidic compared with the cytoplasm. This acidic environment is essential for the activity of many lysosomal acid hydrolases. V-type H⁺-ATPase pumps protons into the lysosome using energy from ATP, helping maintain the required acidic conditions.
How are lysosomes formed?
Lysosome formation involves the rough endoplasmic reticulum, Golgi apparatus, and endosomal system. Lysosomal enzymes are synthesized in the rough ER and processed in the Golgi. Many receive a mannose-6-phosphate tag that helps target them to endosomal compartments. Endosome maturation and trafficking then contribute to the formation of functional lysosomal compartments.
What are the types of lysosomes?
Traditional cell biology describes primary lysosomes, secondary lysosomes, autolysosomes, and residual bodies. Primary lysosomes contain newly delivered lysosomal enzymes, while secondary lysosomes actively digest materials. Autolysosomes form during autophagy when autophagic compartments fuse with lysosomal compartments. Residual bodies contain material remaining after incomplete digestion. Modern biology views lysosomes as dynamic and heterogeneous compartments.
What is the relationship between lysosomes and autophagy?
Lysosomes provide the degradative machinery required for autophagy. During macroautophagy, damaged organelles or cytoplasmic material are enclosed within autophagosomes. These compartments subsequently fuse with lysosomal compartments, forming autolysosomes where the contents are degraded. The resulting molecules, such as amino acids and fatty acids, can be recycled by the cell.
Are lysosomes present in plant cells?
Plant cells generally do not contain typical animal-cell lysosomes. Instead, lytic vacuoles and the central vacuole perform many comparable degradative and recycling functions. These vacuolar compartments contain hydrolytic enzymes and participate in degradation, autophagy-related processes, storage, and cellular homeostasis. Therefore, it is more accurate to describe plant cells as having lysosome-like vacuolar functions.
What happens when lysosomes malfunction?
When lysosomes malfunction, cells may be unable to properly degrade or recycle certain cellular materials. Undegraded substances can accumulate inside lysosomes and interfere with normal cellular functions. Depending on the affected protein or pathway, lysosomal dysfunction can damage tissues and organs and may cause lysosomal storage diseases, neurodegeneration, metabolic abnormalities, or other cellular disorders.
What are lysosomal storage diseases?
Lysosomal storage diseases are a group of inherited metabolic disorders caused by defects affecting lysosomal enzymes, proteins, transporters, or related pathways. These defects prevent normal degradation or trafficking of specific substances, causing them to accumulate inside cells. Examples include Gaucher disease, Tay-Sachs disease, Pompe disease, Fabry disease, and several mucopolysaccharidoses.
What is the difference between a primary and secondary lysosome?
A primary lysosome is traditionally described as a newly formed lysosomal compartment containing hydrolytic enzymes but little material undergoing digestion. A secondary lysosome forms when a lysosomal compartment fuses with a vesicle or compartment containing material to be degraded. Secondary lysosomes therefore represent actively digesting compartments. Modern cell biology emphasizes that lysosomes are dynamic rather than rigid categories.
How many lysosomes are in a cell?
The number of lysosomes varies depending on the cell type, size, and physiological condition. Many animal cells contain hundreds to thousands of lysosomal compartments, rather than a fixed number. Cells with high degradative activity, such as macrophages, may contain more lysosomal compartments. Lysosomes are dynamic organelles whose number and activity can change according to cellular needs.
How are lysosomes formed?
Lysosomes are formed through the coordinated activity of the rough endoplasmic reticulum, Golgi apparatus, and endosomal system. Lysosomal enzymes are synthesized in the rough ER and processed in the Golgi. Many enzymes receive a mannose-6-phosphate tag that helps direct them to endosomes. Endosomal maturation and trafficking then produce functional lysosomal compartments.
How does a lysosome work?
A lysosome works by using acid hydrolases to digest cellular materials inside an acidic lumen. V-type H⁺-ATPase pumps protons into the lysosome, maintaining a pH of about 4.5–5.0. Materials delivered through endocytosis, phagocytosis, or autophagy are broken down, while useful products are transported out for reuse by the cell.
How does a lysosome recycle materials in a cell?
Lysosomes recycle materials by breaking down proteins, lipids, carbohydrates, nucleic acids, and damaged cellular components into smaller molecules. These products include amino acids, sugars, fatty acids, and other metabolites, which can leave the lysosomal compartment and return to cellular metabolic pathways. This recycling helps cells conserve nutrients and maintain cellular homeostasis.
How do lysosomes break down materials?
Lysosomes break down materials using acid hydrolases, which are digestive enzymes that work efficiently in acidic conditions. Proteases digest proteins, lipases break down lipids, nucleases degrade nucleic acids, and glycosidases act on carbohydrates and glycoconjugates. Materials enter lysosomal compartments through endocytosis, phagocytosis, or autophagy and are progressively degraded into reusable components.
How do lysosomes help white blood cells?
Lysosomes help white blood cells destroy and digest pathogens and cellular debris. In phagocytic cells such as macrophages and neutrophils, lysosomal enzymes are delivered to compartments containing engulfed bacteria or other particles. The acidic environment and digestive enzymes help break down these materials. Lysosomes also support immune-cell recycling, processing, and cellular defense.
Conclusion
Lysosomes are essential degradative and recycling organelles that help maintain cellular health and homeostasis.
Their single membrane encloses an acidic lumen containing numerous acid hydrolases, which break down proteins, lipids, carbohydrates, nucleic acids, and other cellular materials.
The acidic environment, maintained mainly by V-type H⁺-ATPase, allows these enzymes to function efficiently.
Beyond intracellular digestion, lysosomes participate in autophagy, endocytosis, phagocytosis, recycling, nutrient regulation, and cellular quality control.
They also work closely with other components of the endomembrane system to process and recycle cellular materials.
When lysosomal enzymes, transport proteins, or related pathways are defective, undegraded substances can accumulate and cause lysosomal storage diseases.
Understanding lysosome structure and function is therefore important for cell biology, biochemistry, biotechnology, medicine, and competitive examinations such as NEET and CSIR NET.
A clear understanding of these organelles also provides a strong foundation for studying cellular degradation, recycling, and metabolic disorders.
Discover more from Go Life Science
Subscribe to get the latest posts sent to your email.